How Can You Verify a Blood Pressure Monitor Supplier’s CE Documents?
A CE logo or a certificate scan does not establish that a blood pressure monitor can legally be placed on the EU market. Reliable verification requires a consistent chain of evidence: the manufacturer’s Declaration of Conformity, the applicable conformity assessment route, any required notified body certificates, and documentation linking that evidence to the exact device being supplied.
The decisive distinction is between documents that support regulatory conformity and documents that merely report testing or quality-system certification. An authentic document can still be insufficient: it may cover another manufacturer, an unrelated device family, or a conformity assessment route that does not fit the monitor’s intended purpose.
Compare the documents by what they actually prove
There is no single universal “CE certificate” that proves every aspect of a blood pressure monitor’s compliance. Under Regulation (EU) 2017/745, the Medical Device Regulation (MDR), the legal manufacturer declares conformity, while a notified body assesses the elements required by the applicable route.
A package containing an ISO 13485 certificate and electromagnetic compatibility reports may demonstrate useful work, but it leaves a critical gap if the medical-device declaration and required notified body evidence are missing. Conversely, a notified body certificate should not replace checks of the actual model, labeling, and manufacturer identity.
The monitor’s classification determines the evidence required
Medical-device classification follows intended purpose and the applicable classification rules, not the product’s price, sales channel, or description as a “home-use” device.
An electronic blood pressure monitor is an active device. Under MDR Annex VIII, Rule 10, active devices intended for diagnosis and monitoring of physiological processes generally fall within Class IIa. Devices specifically intended to monitor vital physiological parameters where variations could result in immediate danger can fall within Class IIb. A routine home monitor and a device intended for critical-care monitoring therefore should not automatically be assessed as equivalent.
A non-active mechanical sphygmomanometer can follow a different route. Where classified as Class I with a measuring function, commonly called Class Im, notified body involvement is limited to the aspects relating to metrological requirements. This differs substantially from the assessment applicable to an active Class IIa monitor.
Ask for the manufacturer’s classification rationale, including the rule applied and the intended purpose appearing in the instructions for use. A claim that an electronic monitor is “Class I, self-declared” deserves particular scrutiny. Likewise, evidence for a mechanical measuring device cannot be assumed to cover an electronic product carrying a similar brand name.
For devices requiring notified body involvement, the CE marking is accompanied by the relevant body’s four-digit identification number. That number identifies the body involved in the applicable conformity assessment; it is not a product approval number.
Match the declaration to the product, not just the supplier
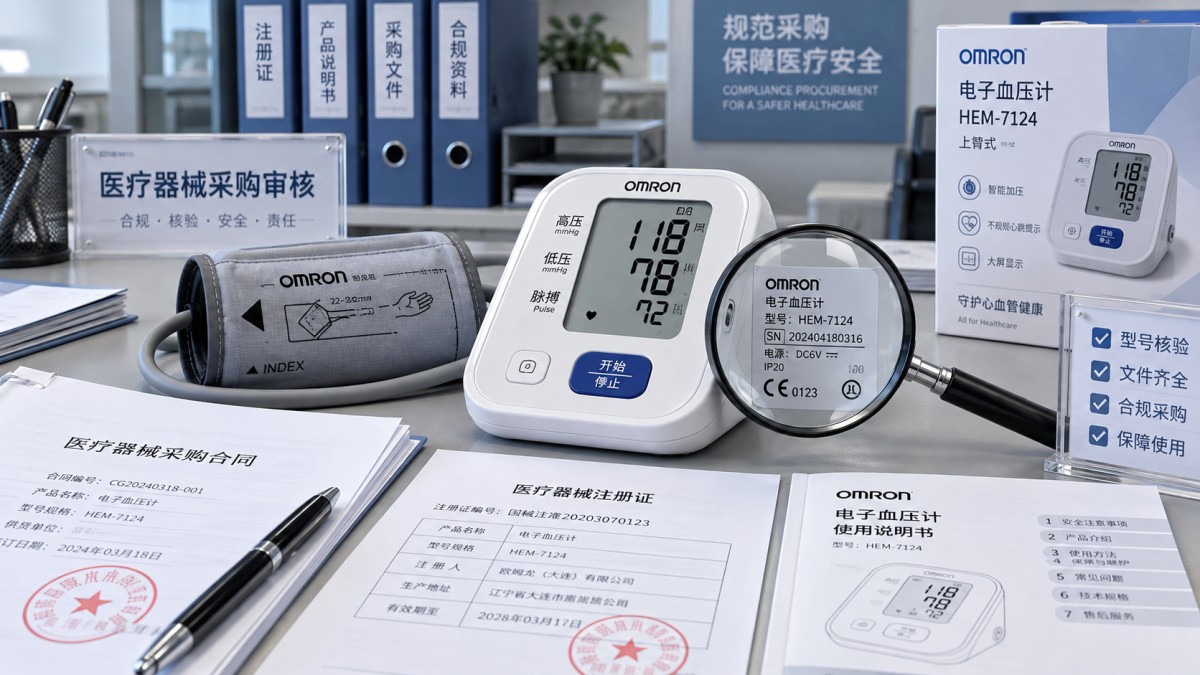
The EU Declaration of Conformity should identify the legal manufacturer and the device clearly enough to establish what is being declared. Under MDR Annex IV, required information includes the manufacturer’s details, a statement of sole responsibility, device identification, risk class, applicable legislation, and relevant notified body information where applicable. The Basic UDI-DI is also part of the MDR declaration requirements.
Obtain the complete, signed declaration rather than a cropped image. Check its issue details, authorized signatory, and all referenced annexes. If device models appear in an attachment, that attachment is part of the verification package, not an optional supplement.
The supplier’s trading name may differ legitimately from the legal manufacturer’s name. What matters is whether the legal manufacturer identified in the declaration matches the manufacturer on the device label and instructions, and whether the commercial relationship explains the supplier’s role.
Private labeling requires closer examination. Selling a device under another company’s name or trademark can create manufacturer obligations under MDR Article 16, subject to the conditions and exceptions specified there. A factory certificate does not automatically cover a separately branded device simply because the hardware appears identical.
Model differences also need an explanation. A declaration for “BP-200 Series” may cover “BP-200A,” but that relationship should be supported by controlled manufacturer documentation. A reseller’s email saying “all models use the same certificate” is not equivalent evidence. Different measurement algorithms, cuff configurations, intended populations, or intended uses can affect coverage.
Use NANDO to check designation—and the issuer to check authenticity
Two separate questions must be answered: is the organization authorized to perform the relevant assessment, and did it genuinely issue the certificate presented?
The European Commission’s NANDO notified body information system addresses designation. Search using the four-digit number and confirm the organization’s identity, the applicable legislation, and the scope of its designation. A designation under another product regulation, or under the former Medical Devices Directive, is not evidence of an MDR designation.
Scope matters as much as the organization’s name. A body’s authority is defined by its designation; being listed somewhere in NANDO does not authorize it to assess every medical device or perform every conformity assessment activity.
NANDO does not authenticate an individual certificate. For that, use the issuing body’s official certificate-verification service or contact details obtained independently from its official website. Avoid relying exclusively on a supplier-provided email address, QR code, or verification link.
Where confirmation is needed, provide the certificate number, manufacturer’s name, issue date, and full document. Request confirmation of authenticity, current status, scope, and any relevant restrictions or replacement certificates. A genuine certificate may have been suspended, withdrawn, superseded, or narrowed after its original issue.
An MDR quality management system certificate may describe a device category rather than list every commercial model. Absence of a model number on its face is therefore not automatically disqualifying. However, the declaration, certificate scope, and supporting manufacturer records must still establish that the offered monitor falls within the assessed system and relevant device scope. Missing certificate schedules can prevent that conclusion.
Compare MDR certification with legacy MDD evidence carefully
Documents referring to Directive 93/42/EEC, the former Medical Devices Directive (MDD), require a different analysis from documents issued under the MDR. They should not be rejected solely because they reference the older legislation, but they should not be accepted solely because a supplier claims an automatic extension.
Regulation (EU) 2023/607 extended transitional arrangements for qualifying legacy devices. For eligible Class IIa devices, the transition can extend to 31 December 2028, provided the relevant conditions are met. Eligibility depends on more than device class: conditions include continued compliance with the former directive, no significant changes in design or intended purpose, and fulfillment of specified MDR transition requirements.
The transition framework included deadlines for implementing an MDR-compliant quality management system, submitting a formal application to a notified body, and entering into a written agreement. Consequently, an old certificate’s printed expiry date does not by itself determine whether the device remains eligible. Equally, a supplier cannot establish eligibility merely by attaching a copy of the amending regulation.
For legacy documentation, request the declaration under the applicable directive, the original certificate and schedules, and evidence supporting continued transitional eligibility. This can include the manufacturer’s transition statement and a notified body confirmation letter identifying the relevant devices and certification arrangements.
The important comparison is between a documented transition position and an unsupported assertion that “all CE certificates were extended.” Where scope or eligibility remains unclear, resolve it with the manufacturer and relevant notified body before treating the documentation as acceptable.
Technical reports should support the same device configuration
Regulatory documents establish the legal conformity position; technical evidence helps assess whether that position credibly relates to the supplied monitor. These checks overlap, but they are not interchangeable.
For an automated non-invasive blood pressure monitor, relevant evidence can include electrical safety and essential performance testing, electromagnetic compatibility testing, and clinical investigation of measurement accuracy. Standards such as IEC 80601-2-30 and ISO 81060-2 may be relevant, depending on the device and claimed conformity approach.
A standard number on a brochure is not evidence of testing. Review the report’s model identification, tested configuration, edition, amendments, results, and issuing laboratory. Where a report covers a related model, request the technical justification explaining why its results apply to the supplied version.
Do not confuse a pressure-sensor calibration check with clinical validation of blood pressure measurement accuracy. An oscillometric monitor estimates blood pressure using signals and an algorithm; demonstrating accurate pressure measurement alone does not establish that the complete device produces clinically accurate readings.
Claims covering special populations, such as children or pregnancy, require evidence appropriate to those intended uses. Cuff size and configuration also matter. A report for one cuff configuration should not be assumed to substantiate every accessory offered with the product.
Standards provide routes for demonstrating particular requirements, but not every published standard is harmonized under the MDR. If a declaration claims harmonized standards, check the relevant references in the EU’s published lists. Standard conformity does not eliminate the need for the applicable medical-device assessment.
Separate explainable gaps from reasons to stop acceptance
Some discrepancies are administrative: an omitted annex, an outdated address following a documented company change, or a model-family designation requiring clarification. They still need resolution, but they differ from evidence that points to the wrong regulatory route or an unrelated manufacturer.
Stronger warning signs include a certificate number the issuer cannot authenticate, altered pages, a notified body number belonging to another organization, or a declaration that cites only general electrical legislation for a medical blood pressure monitor. Refusal to supply the complete declaration or explain model coverage is also materially different from a short delay in obtaining an archived schedule.
A practical acceptance record should retain the exact declaration, certificate versions, independent verification results, classification rationale, and model-mapping evidence. Record the label and instructions reviewed against them. Before shipment release, confirm that the supplied branding, model, manufacturer details, and CE marking remain consistent with that record.
No single database search or certificate scan proves the entire compliance position. Acceptance is defensible when the applicable route is credible, the required documents are authentic and current, and the evidence consistently identifies the device being supplied. If that chain breaks, additional certificates unrelated to the missing link do not repair it.
The primary legal references are Regulation (EU) 2017/745, particularly its declaration, classification, and conformity assessment provisions, and Regulation (EU) 2023/607 for the amended transitional arrangements. These should take precedence over supplier summaries when determining the applicable requirements.
 Can you trust flexible packaging market intelligence forecasts?Can you trust flexible packaging market intelligence forecasts? Learn how to verify sources, challenge assumptions, and assess demand before making costly investment or sourcing decisions.
 When is supply chain disruption management consulting worth the cost?When is supply chain disruption management consulting worth the cost? Learn how to compare fees, avoidable losses, and recovery options to make a confident investment decision.
 How to use macroeconomic analysis to forecast industrial demandApply an industrial macro analysis methodology to forecast demand, interpret economic signals, and plan inventory and capacity. Discover practical steps for smarter business decisions.
 How to assess Singapore suppliers with global supply chain intelligenceExplore international supply chain intelligence Singapore buyers can use to verify suppliers, assess capabilities and delivery risks, and make confident sourcing decisions.
 Which macroeconomic indicators help forecast U.S. industrial demand?Explore industrial macro analysis USA decision-makers can use to forecast demand. Track new orders, inventories, and credit signals to sharpen procurement and capacity decisions.
 Can a food processing manufacturer meet your small-batch needs?Can a food processing manufacturer meet your small-batch needs? Discover how to assess minimum orders, costs, packaging, and safety to choose the right partner for your launch.
 How to verify sourcing intelligence data before selecting suppliersVerify sourcing intelligence data before choosing suppliers. Learn how to check company records, certifications, and capacity to reduce procurement risk and build a reliable shortlist.
 How to use supplier risk analysis to spot financial warning signsUse supplier risk analysis to spot cash flow pressure, debt risks, and delivery warning signs. Learn practical steps to reduce exposure and protect supply continuity.
 When dual sourcing makes sense in a sourcing strategy for manufacturersSourcing strategy for manufacturers: learn when dual sourcing strengthens supply continuity, reduces disruption risk, and protects production without adding unnecessary complexity.
 How bulk commodities pricing affects long-term supply contractsBulk commodities pricing shapes long-term supply contracts through index rules, freight, quality, volume, and currency risk. Explore strategies for resilient costs and supply security.
 Which global manufacturing trends by sector matter most for sourcing plans?Global manufacturing trends by sector: identify sourcing risks, supplier resilience, compliance demands, and cost drivers to build smarter, more reliable procurement plans.
 What industrial trade intelligence pricing should include before you subscribeIndustrial trade intelligence pricing explained: compare coverage, data quality, access rights, and workflow value before choosing a subscription that supports smarter trade decisions.
 How an industrial trade intelligence dashboard can identify new supplier marketsDiscover how an industrial trade intelligence dashboard helps identify new supplier markets by comparing capacity, landed cost, logistics resilience, and compliance risk.
 Which global market entry barriers should exporters assess first?Global market entry barriers exporters should assess first: tariffs, compliance, buyer qualification, logistics, and payment risk. Build a practical strategy for confident market entry.
 How trade data aggregation analytics can reveal emerging supplier marketsTrade data aggregation analytics reveals emerging supplier markets by connecting trade flows, capacity signals, logistics, and compliance insights for smarter sourcing decisions.
 When an export market analysis platform can validate demand in a new marketExport market analysis platform insights help validate new-market demand through buyer fit, regulations, logistics, competition, and profitable entry conditions.
 What is driving the beauty and personal care market in the USA?Beauty and personal care in USA is driven by targeted solutions, selective premiumization, digital discovery, clean-label trust, diverse needs, and compliance. Explore key growth opportunities.
 How trade flow monitoring reports reveal changing regional demandTrade flow monitoring reports reveal changing regional demand through import volumes, sourcing shifts, product mix, logistics signals, and regulatory trends. Explore actionable market insights.
 How international market intelligence for investors reduces market-entry riskInternational market intelligence for investors reduces market-entry risk by validating demand, regulations, logistics, and competition before capital is committed.
 How international sourcing intelligence helps identify supplier risk earlyInternational sourcing intelligence helps businesses spot supplier risks early, protect supply continuity, and make smarter global procurement decisions.
 How trade data aggregation analytics reveals emerging supplier marketsTrade data aggregation analytics reveals emerging supplier markets through export trends, input flows, logistics, and regulatory signals. Discover smarter sourcing insights.
Global Trade Insights & Industry
Our mission is to empower global exporters and importers with data-driven insights that foster strategic growth.
Search News
Popular Tags
Industry Overview
The global commercial kitchen equipment market is projected to reach $112 billion by 2027. Driven by urbanization, the rise of e-commerce food delivery, and strict hygiene regulations.